Patient Information
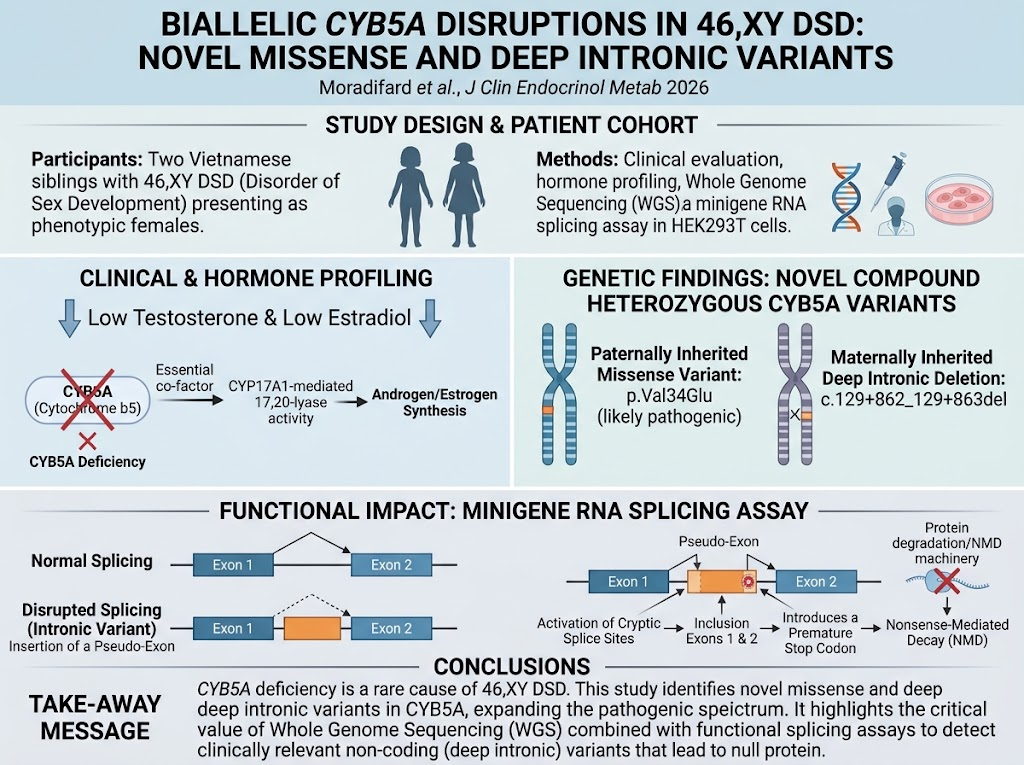
Two siblings from a Vietnamese family, genetically male (46,XY karyotype), presented clinically at pediatric age with a female phenotype. Both were phenotypic females at birth with no ambiguous genitalia noted. The family history was notable for consanguinity not reported; no prior similar cases in the family were documented. Their main clinical concern was disorders of sex development (DSD) detected during pediatric evaluations.
Diagnosis
Clinical evaluation included hormone profiling revealing low serum testosterone and estradiol levels inconsistent with their 46,XY genotype, indicating impaired steroidogenesis. The atypical phenotype prompted molecular investigations. Whole genome sequencing (WGS) of DNA extracted from both siblings identified compound heterozygous variants in the CYB5A gene: one missense variant, p.Val34Glu, inherited from the father, and a deep intronic deletion, c.129+862_129+863del, inherited maternally.
The intronic deletion was predicted using SpliceAI to activate cryptic splice sites, potentially leading to aberrant splicing. Functional validation was performed using a minigene RNA splicing assay in HEK293T cells, which confirmed activation of these cryptic sites and inclusion of a pseudo-exon between exons 1 and 2 of CYB5A. This aberrant insertion introduced a premature termination codon likely causing nonsense-mediated decay (NMD) of the transcript, resulting in loss of functional cytochrome b5 protein.
According to ACMG/AMP 2015 guidelines, both variants were classified as likely pathogenic based on their predicted and experimentally confirmed deleterious effects. The genetic diagnosis of CYB5A deficiency was thus established, explaining the 46,XY DSD phenotype.
Differential Diagnosis
Differential diagnoses considered included other etiologies of 46,XY DSD such as androgen insensitivity syndrome (AIS), 5-alpha-reductase deficiency, and steroidogenic enzyme deficiencies including 17-beta hydroxysteroid dehydrogenase deficiency or CYP17A1 mutations. AIS was less likely due to hormonal profile showing low androgens rather than receptor resistance. 5-alpha-reductase deficiency usually presents with ambiguous genitalia but elevated testosterone levels, and CYP17A1 mutations generally affect both 17-hydroxylase and 17,20-lyase activities differently. The hormone pattern and genetic findings specifically implicate CYB5A deficiency impacting CYP17A1 17,20-lyase activity.
Treatment and Management
Management focused on addressing the hormonal insufficiency and potential psychosocial considerations due to the discordance between genotype and phenotype. Hormone replacement therapy was considered to induce secondary sexual characteristics aligned with the patient’s phenotypic sex of upbringing. Multidisciplinary care involving endocrinology, genetics, psychology, and supportive counseling was recommended. Genetic counseling provided the family with information regarding the recessive inheritance, implications for future pregnancies, and available reproductive options.
Outcome and Prognosis
Follow-up data are limited; however, with early diagnosis, tailored hormone replacement and support, patients with CYB5A-related 46,XY DSD can achieve appropriate pubertal development and improved psychosocial outcomes. Long-term prognosis depends on hormone management and adaptation to individual gender identity.
Discussion
Cytochrome b5 (encoded by CYB5A) is a critical cofactor for CYP17A1, an enzyme essential for sex steroid synthesis, particularly the 17,20-lyase activity needed to produce androgens. CYB5A deficiency leads to isolated 17,20-lyase deficiency, a rare cause of 46,XY DSD resulting in undervirilized or phenotypic female genitalia despite male karyotype.
This case highlights several novel insights. Firstly, the two siblings exhibited compound heterozygosity for a novel missense variant and a deep intronic deletion disrupting splicing. Deep intronic variants are often missed by traditional gene panels but can produce significant functional effects, which here was proven by minigene splicing assays confirming pseudo-exon inclusion and transcript disruption. This underscores the value of whole genome sequencing combined with functional validation in diagnosing elusive genetic causes of DSD.
Secondly, this finding expands the mutational spectrum of CYB5A-related DSDs, previously limited to a few reported variants, and provides further genotype-phenotype correlation. The lack of testosterone and estradiol aligns with impaired steroidogenesis at the 17,20-lyase step due to absent functional cytochrome b5.
Clinically, this case supports including CYB5A sequencing in the diagnostic workup for 46,XY DSD patients with low androgens and normal cortisol pathways. It also stresses that non-coding variants must be considered, particularly in patients where single coding region mutations are not detected.
Finally, early molecular diagnosis facilitates appropriate clinical management, prognosis assessment, and genetic counseling, ultimately improving patient care.
References
1. Moradifard S, Le TNU, Ha NT, et al. Biallelic CYB5A disruptions in 46, XY DSD: Novel missense and deep intronic variants. The Journal of Clinical Endocrinology and Metabolism. 2026; Oct 07. PMID: 42839303.
2. Flück CE, Pandey AV. Impact of CYP17A1 deficiency on steroidogenesis characterized by altered 17,20-lyase activity. Horm Res Paediatr. 2020;93(1):21-32.
3. Blau JE, Auchus RJ. The spectrum of CYP17A1 deficiency: steroids, structure, and function. J Steroid Biochem Mol Biol. 2021;216:105926.
4. Richards S, Aziz N, Bale S, et al. Standards and guidelines for the interpretation of sequence variants: ACMG/AMP. Genet Med. 2015;17(5):405-424.
5. Soemedi R, Cygan KJ, Rhine CL, et al. A deep intronic mutation causes pseudoexon activation resulting in congenital adrenal hyperplasia. Hum Mutat. 2017;38(3):289-295.
This case report underlines the importance of integrating genetic technologies with functional assays to uncover pathogenic mechanisms in rare endocrine disorders, enabling precise diagnosis and targeted management of 46,XY DSD.